2027 Mass Analytica Workshops
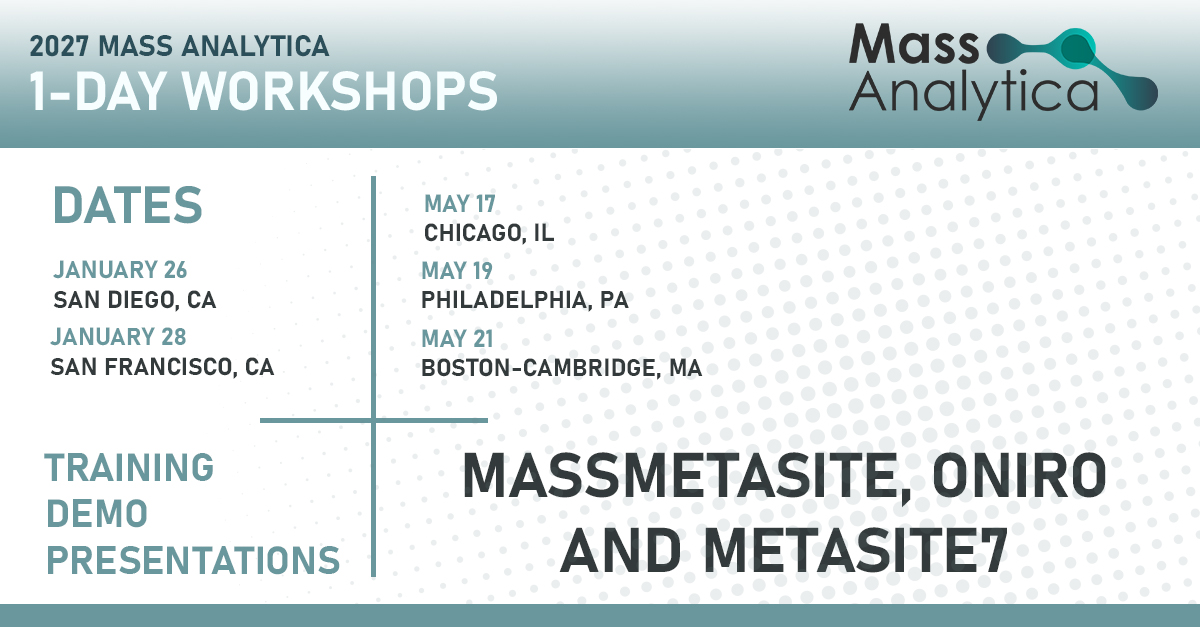
We are pleased to announce the Mass Analytica 2027 Workshop Series, with five in-person sessions across the United States.
REVENG: 3D-DRIVEN REVERSE ENGINEERING OF METABOLIC BIOLOGICAL PATHWAYS
REVENG: 3D-DRIVEN REVERSE ENGINEERING OF METABOLIC BIOLOGICAL PATHWAYS
September 2026, 25th EuroQsar
Tommaso Palomba1,2, Savannah Mason1, Paula Cifuentes1, Ismael Zamora1, Gabriele Cruciani2
1Mass Analytica, S.L., Sant Cugat del Vallés, Spain; 2Molecular Discovery Ltd, Kinetic Business Centre, Borehamwood, UK
Abstract
Elucidating the specific enzymes responsible for metabolite formation is a cornerstone of drug discovery; it is essential for mapping metabolic pathways, assessing drug-drug interactions, driving lead optimization to enhance metabolic properties, and mitigating toxicity risk. While liquid chromatography-mass spectrometry (LC-MS/MS) excels at structural Metabolite Identification (MetID), experimental reaction phenotyping remains a major resource-intensive bottleneck. To bridge this gap, we present RevEng, an innovative computational workflow integrated within MetaSite 71. Taking the parent drug and a target metabolite (for example, an experimentally detected LC-MS/MS metabolite) as inputs, RevEng structurally reverse-engineers the metabolic space to reconstruct all potential metabolic pathways, including complex multi-step biotransformations, and assigns a probability score to the enzymes involved in each step. This inverse pathway prediction relies entirely on structure-based 3D docking and intrinsic chemical reactivity, using dedicated enzyme-specific predictive models for 23 Phase I (CYP and non-CYP) and 20 Phase II major human drug-metabolizing enzymes2,3. These models are interaction- and reaction-based and are therefore fully independent of training-set-derived substrate knowledge. Enzyme active sites are characterized using GRID Molecular Interaction Fields (MIFs)4, while the docking procedure is specifically designed to mimic the biological and chemical determinants of each enzymatic reaction by evaluating substrate–residue interactions within the catalytic site. The resulting interaction energy, combined with an assessment of the molecule’s intrinsic reactivity toward a specific biotransformation, enables prediction of the most likely site of metabolism, related metabolites structures and the likelihood of metabolism by the enzyme under consideration. The workflow was rigorously validated against proprietary in-house experimental datasets, and the predicted inverse pathways were benchmarked against established literature data for diverse chemical entities. The 3D-docking engine successfully unraveled complex metabolic fates, accurately backtracking both single-step biotransformations and sequential, multi-step cascades (such as Phase I functionalization followed by Phase II conjugation). Beyond pathway elucidation, the prediction of the enzymes responsible for the formation of observed metabolites can directly drive experimental design, enabling the detection of metabolites using targeted enzymatic assays rather than complex biological matrices, thus leading to a better characterization of the metabolic profile of a drug. Furthermore, these predictions provide invaluable guidance for targeted metabolite synthesis and for anticipating potential toxicity liabilities. In conclusion, RevEng, in combination with the other advanced tools integrated within MetaSite 7, successfully bridges analytical chemistry and mechanism-based 3D docking, offering a chemically accurate and structurally-driven solution for automated reaction phenotyping and intelligent experimental design in early drug discovery.
- Isoherranen, N., & Zanger, U. M. (2025). Cytochrome P450 enzymes in drug metabolism and precision medicine: Current challenges and future directions. ACS Pharmacology & Translational Science, 8(2), 114-131.
- Cruciani, G., Carosati, E., De Boeck, B., Ethirajulu, K., Mackie, C., Howe, T., & Vianello, R. (2005). MetaSite: Understanding Metabolism in Human Cytochromes from the Perspective of the Chemist. Journal of Medicinal Chemistry, 48(22), 6970-6979.
- Di, L., & Obach, R. S. (2024). The evolving landscape of non-CYP enzymes in drug metabolism: Current status and clinical relevance. Drug Metabolism Reviews, 56(2), 145-168.
- Artese, A., Cross, S., Costa, G., Distinto, S., Parrotta, L., Alcaro, S., Ortuso, F., & Cruciani, G. (2013). Molecular interaction fields in drug discovery: recent advances and future perspectives. WIREs Computational Molecular Science, 3(6), 594-613.
You must be logged in to access this content. Not yet registered? Create a new account
PREDICTION OF PHASE I AND PHASE II METABOLISM
PREDICTION OF PHASE I AND PHASE II METABOLISM
June 2026, ISSX 16th European Meeting
Tommaso Palomba1,2, Ludovico Venturi1,2, Massimo Baroni1, Paolo Benedetti1, Gabriele Cruciani1,2,3
1Mass Analytica, S.L. Sant Cugat del Vallès, Spain; 2Molecular Discovery Ltd, Kinetic Business Centre, Borehamwood, UK; 3Department of Chemistry, Biology, and Biotechnology, University of Perugia, Italy
Abstract
Many drugs are metabolized in the human body to form metabolites for elimination. This metabolism is categorized into two classes of reactions. Phase I reactions are mediated by Cytochrome P450 enzymes, in addition to approximately thirty other enzymes. An additional twenty other enzymes are involved in Phase II reactions, which often result in conjugation products.
While the cytochrome P450 enzymes have long been studied, there is a growing need to better understand the role of non-CYP enzymes in drug metabolism. To facilitate this, we developed MetaSite 7 to provide the prediction of drug metabolism using 23 non-CYP Phase I enzymes including eight families and their related isoforms. Twenty enzymes, including 12 different families, responsible for Phase II reactions were also considered.
The prediction determines the probability of a molecule to be a substrate for each enzyme by identifying reactive atoms in the molecule structure. In MetaSite 7, we used the enzyme’s 3D structure to identify reactive atoms of the molecule which may be exposed to the cofactor or active binding site of the enzyme. The GRID force field is used to calculate flexible interaction fields between the catalytic residues of the enzyme and the molecule.
Multiple docking poses are considered, and the energy contributions are summed. The poses are ranked, and the lowest energy pose is used to generate a [1,2,3]This prediction can be repeated for each of the enzymes in MetaSite 7, resulting in a more comprehensive view of the site of metabolism (SOM) for both Phase I and Phase II enzymes.
Furthermore, the predictions may be combined by subjecting the potential first-generation metabolites to further predictions, allowing for a more complete view of second-generation metabolites.
In conclusion, we utilized the 3D structures of the different CYP isoforms and 43 non-CYP enzymes to predict both Phase I and Phase II metabolism, along with the probability of a drug molecule to be a substrate for each enzyme. These predictions provide an increased understanding of the potential metabolic pathway in the human body.
You must be logged in to access this content. Not yet registered? Create a new account
GRAPHORMER BASED PREDICTION OF PEPTIDE METABOLIC STABILITY AND PHARMACOKINETIC RELATED PROPERTIES
GRAPHORMER BASED PREDICTION OF PEPTIDE METABOLIC STABILITY AND PHARMACOKINETIC RELATED PROPERTIES
June 2026, ISSX 16th European Meeting
Ismael Zamora1, Albert Garriga1, Luca Morettoni1, Paula Cifuentes2,3, Ramon Adàlia2,4
1Mass Analytica, S.L., Sant Cugat del Vallés, Spain; 2Lead Molecular Design, SL, Sant Cugat del Vallès, Spain 3Universitat Pompeu Fabra, Barcelona, 08003, Spain, 4Universitat Autònoma de Barcelona, Cerdanyola del Vallès, Spain
Abstract
Peptides represent a promising class of therapeutic agents due to their high target specificity, favorable safety profiles, and efficient tissue penetration. However, their clinical development remains limited by poor in vivo stability, rapid enzymatic degradation, and short plasma half-lives. Structural flexibility and high solvent accessibility further increase susceptibility to proteolysis, resulting in rapid clearance and reduced therapeutic efficacy. While chemical strategies such as cyclization and incorporation of non-natural amino acids can improve peptide stability, it remains challenging prior to synthesis to identify peptides that would benefit from such modifications based on metabolic stability and pharmacokinetic properties.
Here, we present a Graphormer-based machine learning framework for predicting key properties relevant to peptide metabolic stability, including proteolytic cleavage sites and peptide blood stability, as well as pharmacokinetic properties such as permeability. In addition, we demonstrate the ability of the approach to predict other peptide properties, including solvent accessibility, secondary structure, and post-translational modifications, which are included to assess the model’s capacity to capture structural information relevant for predicting metabolism- and pharmacokinetics-related properties.
The models employ transformer architecture with added mechanisms to encode graph structural information. Importantly, unlike most existing state-of-the-art tools, our approach is not restricted to natural amino acids and supports cyclic peptides. Depending on the target property, the Graphormer architecture was adapted for different downstream tasks, including binary or multiclass classification, ranking, and regression.
The Graphormer site of cleavage ranking model was evaluated under 28 datasets collected from MEROPS, which compiled experimentally verified cleavage sites from proteases involved in peptide-drug degradation. The site-of-cleavage ranking model was evaluated on 28 MEROPS datasets of experimentally verified protease cleavage sites involved in peptide-drug degradation, achieving an average MAP of 0.59 and Precision@1 of 0.47, outperforming ProsperousPlus.¹
The peptide blood stability model, trained on 635 peptides up to 50 amino acids including cyclic and modified peptides, achieved comparable performance to PepMSND², with accuracy 0.84, precision 0.85, recall 0.81, F1 score 0.81, and MCC 0.70.
For permeability, trained and tested on 6,888 cyclic peptides from the CycPeptMP database³, the model achieved R² = 0.77, MAE = 0.36, and MSE = 0.26, showing good generalization to structurally diverse peptides.
For solvent accessibility, formulated as buried versus exposed residues and validated on protein sequences up to 1,000 amino acids, the model achieved recall 0.81 and F1 score 0.78, comparable to existing methods.⁴
For secondary structure prediction evaluated using Jensen-Shannon distance across helix, strand, and coil states, the model achieved the lowest JSD for coil (0.207), outperforming all compared methods⁵, and improved over JPred4 and PSIPRED for helix and strand while narrowing the gap with PEP2D.
Finally, in predicting post-translational modifications across seven PTM types, the binary model outperformed state-of-the-art method⁶ for lysine acetylation, SUMOylation, ubiquitination, and arginine methylation, and achieved comparable performance for the remaining classes, demonstrating robustness across diverse biochemical modifications.
You must be logged in to access this content. Not yet registered? Create a new account